BAM Statistics (Single-End)

Output BAM Header including alignment statistics and parameters given any indexed (BAI) BAM File.
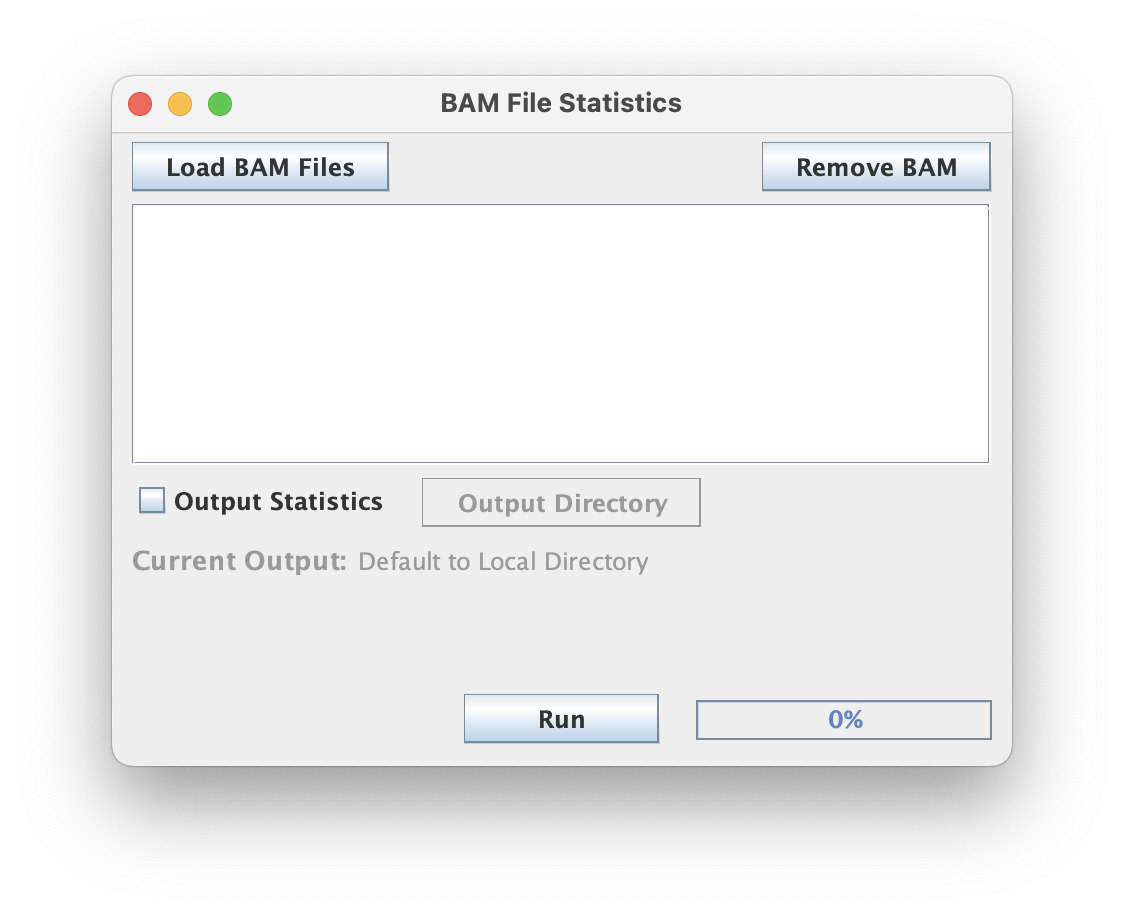
This script processes each input BAM file and tallies up reads per chromosome.
File input (BAM)
This script processes BAM-type files so make sure your input is properly formatted and uses the appropriate .bam extension if you are working from the GUI.
Output statistics (TXT)
By default, the Paired-End Statistics does not record the computed values (often users just want to take a look at the report) so check this box if you wish to save these results. If you do check this box, each input file will have a *_SE-stats.txt file recording the reported statistics.
Even if this output box is not set, note the [logging manager][scriptmanager-logging] will track this run as if this checkbox was selected.
Command Line Interface
Usage:
java -jar ScriptManager.jar bam-statistics se-stat <bamFile>
[-hV] [-o=<output>]
Positional Input
This tool takes a single BAM file for input. As with other tools, this tool requires the BAM file be indexed.
Output Options
| Option | Description |
|---|---|
-o, --output=<output> | Specify output file |