BAM Correlation

Genome-Genome correlations for replicate comparisons given multiple sorted and indexed (BAI) BAM files.
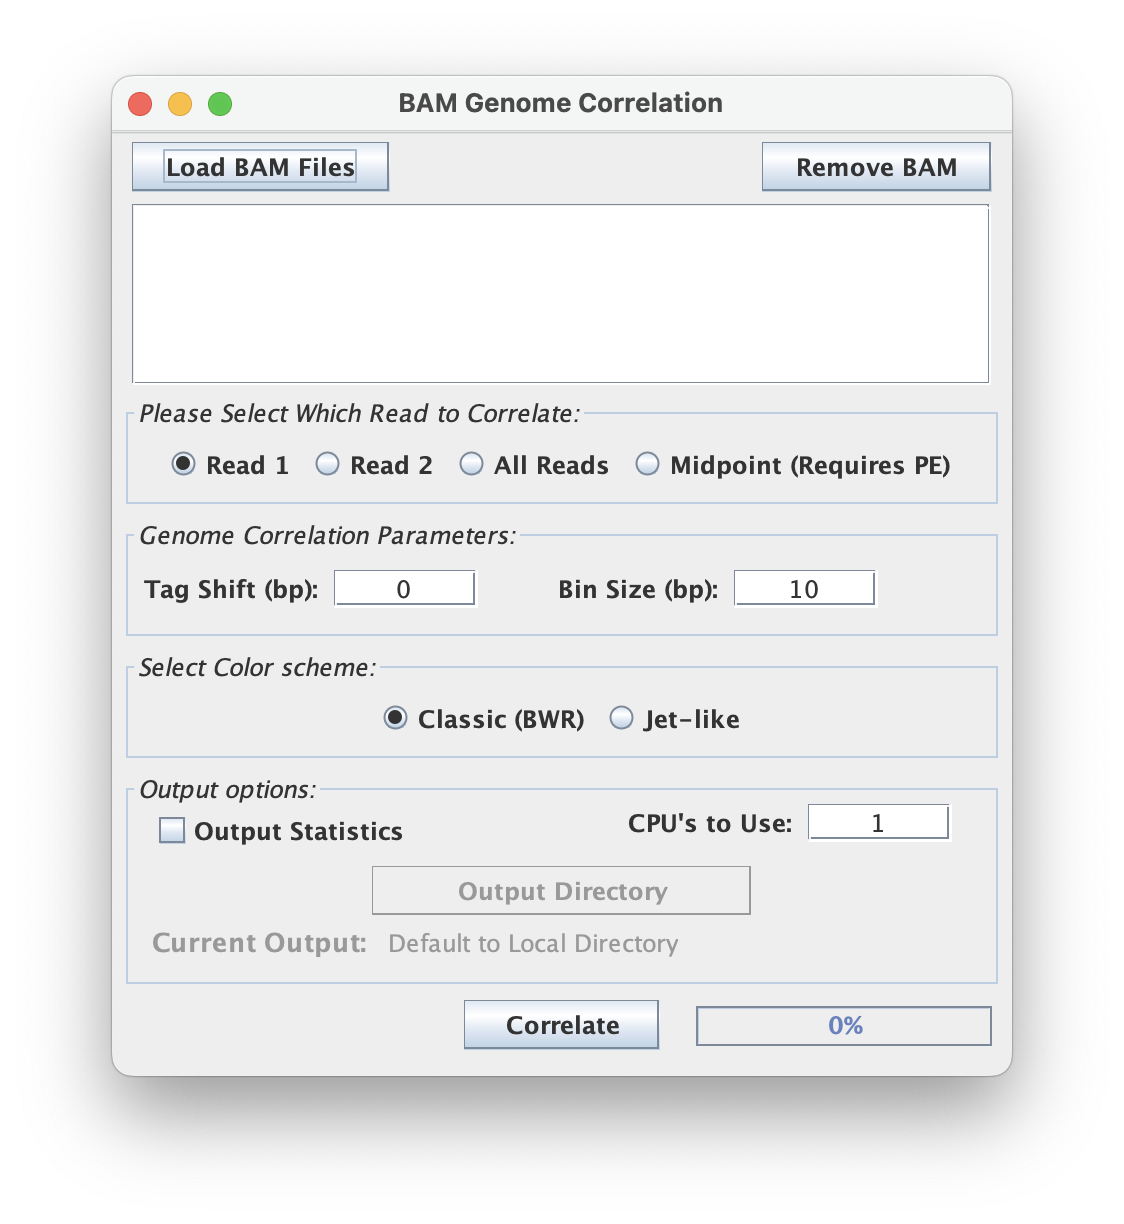
This script processes each input BAM file and peforms a genome-wide correlation for every pair.
File input (BAM)
This script processes BAM-type files so make sure your input is properly formatted and uses the appropriate .bam extension if you are working from the GUI.
Note that unlike most other ScriptManager tools, this script does not process each BAM file independently, but rather computes a relationship between every pair of BAM files provided (Many BAM --> one output as opposed to Each BAM --> one output).
Read Options
This tool has multiple read options to output.
- Read 1: output Read 1
- Read 2: output Read 2
- All Reads: output both reads combined
- Midpoint (Requires PE): output the midpoint between reads
Make sure your BAM input files are [sorted][sort-bam] and [indexed][bam-indexer].
Command Line Interface
Compare a list of BAM files to get a matrix of correlations between them. Outputs both a text file of matrix correlation scores and a heatmap PNG.
Usage:
java -jar ScriptManager.jar bam-statistics bam-correlation
[-1 | -2 | -a | -m] [-fhV] [-b=<binSize>] [--cpu=<cpu>]
[-o=<outputBasename>] [-t=<tagshift>] [<inputFiles>...]
Input Options
| Option | Description |
|---|---|
<inputFiles>... | The BAM file(s) whose statistics we want. |
-f, --files | Input file list of BAM filepaths to correlate (formatted so each path is on its own line) |
Since this tool process a bunch of files together, there are two ways of feeding input files:
(1) You can list them out in the command line tool,
java -jar ScriptManager.jar bam-statistics bam-correlation
bamFile1 bamFile2 ... bamFileX <OPTIONS>
(2) or you can write all the paths for all your files in a single file and pass that as the input using the -f flag
java -jar ScriptManager.jar bam-statistics bam-correlation inputFile -f <OPTIONS>
...where inputFile is listed out line by line:
/path/to/bamFile1
/path/to/bamFile2
...
/path/to/bamFileX
Note that absolute file paths are easier to work with. For relative paths, you'll have to check that they are built with respect to the ScriptManager directory.
Output Options
| Option | Description |
|---|---|
-o, --output=<outputBasename> | Specify output file, default is "correlation_matrix" or the input filename if -f flag used |
| Option | Description |
|---|---|
-t, --tag-shift=<tagshift> | tag shift in bp (default 0) |
-b, --bin-size=<binSize> | bin size in bp (default 10) |
--cpu=<cpu> | CPUs to use (default 1) |
Read Options
| Option | Description |
|---|---|
-1, --read1 | output read 1 (default) |
-2, --read2 | output read 2 |
-a, --all-reads | output combined |
-m, --midpoint | output midpoint (require PE) |